 USA Regulatory Services
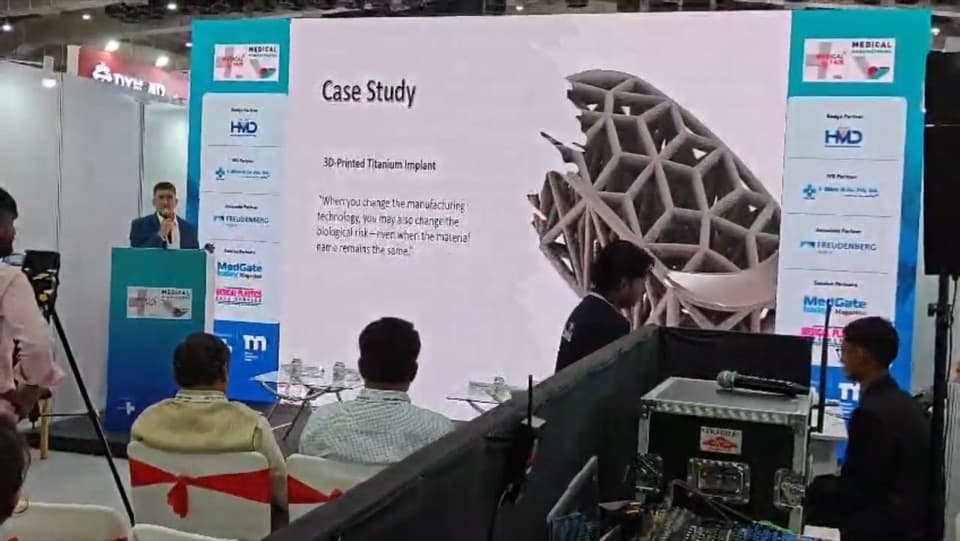
Medreg offers specialized training programs designed to enhance regulatory knowledge and practical understanding of medical device compliance requirements. Our training covers key regulatory frameworks, submission pathways, quality management requirements, and technical documentation applicable to medical devices, with a focus on FDA regulations and other relevant international requirements.
- FDA Medical Device Classification
- Premarket Notifications (510(k)) Submissions
- FDA Establishment Registration and Device Listing
- UDI and GUDID Registration
- QMSR and ISO 13485 Compliance
- Design Control Documentation
- Medical Device Labelling
- Biocompatibility and Biological Safety Evaluation
- Risk Management and Technical Documentation
- Regulatory Reporting and Medical Device Reporting (MDR)
- FDA Q-Submission / Pre - submission
- Device Master Record (DMR) and Device History Record (DHR)
- FDA Inspection Readiness
- Import, Export
- U.S. Agent Requirements
The programs are intended to help medical device manufacturers, regulatory affairs professionals, quality assurance teams, and other industry personnel understand applicable regulatory obligations and develop the skills needed to implement effective compliance practices.
Our training sessions address both the fundamental principles and practical aspects of regulatory compliance, helping participants understand device classification, regulatory pathway selection, premarket submission preparation, quality system implementation, labelling requirements, and biological safety evaluation.